The current medical management of Stickler Syndrome is largely reactive, focusing on mitigating symptoms and preventing complications. Patients rely on corrective lenses for myopia, surgical interventions for cleft palate or joint issues, and prophylactic procedures to guard against retinal detachment. However, the holy grail of inherited retinal disease (IRD) research is to move beyond symptom management and address the root cause of the disorder at the genetic level. While still in the early stages of preclinical research, advances in gene editing and molecular therapeutics are beginning to illuminate potential future pathways for treating Stickler Syndrome.
The Unique Challenge of Collagen Disorders
Developing genetic therapies for Stickler Syndrome presents unique scientific hurdles. Many successful gene therapies currently in use or in advanced clinical trials (such as Luxturna for RPE65-mediated inherited retinal dystrophy) utilize a "gene addition" approach. In these cases, a viral vector delivers a healthy, functional copy of a gene to replace a missing or defective enzyme.
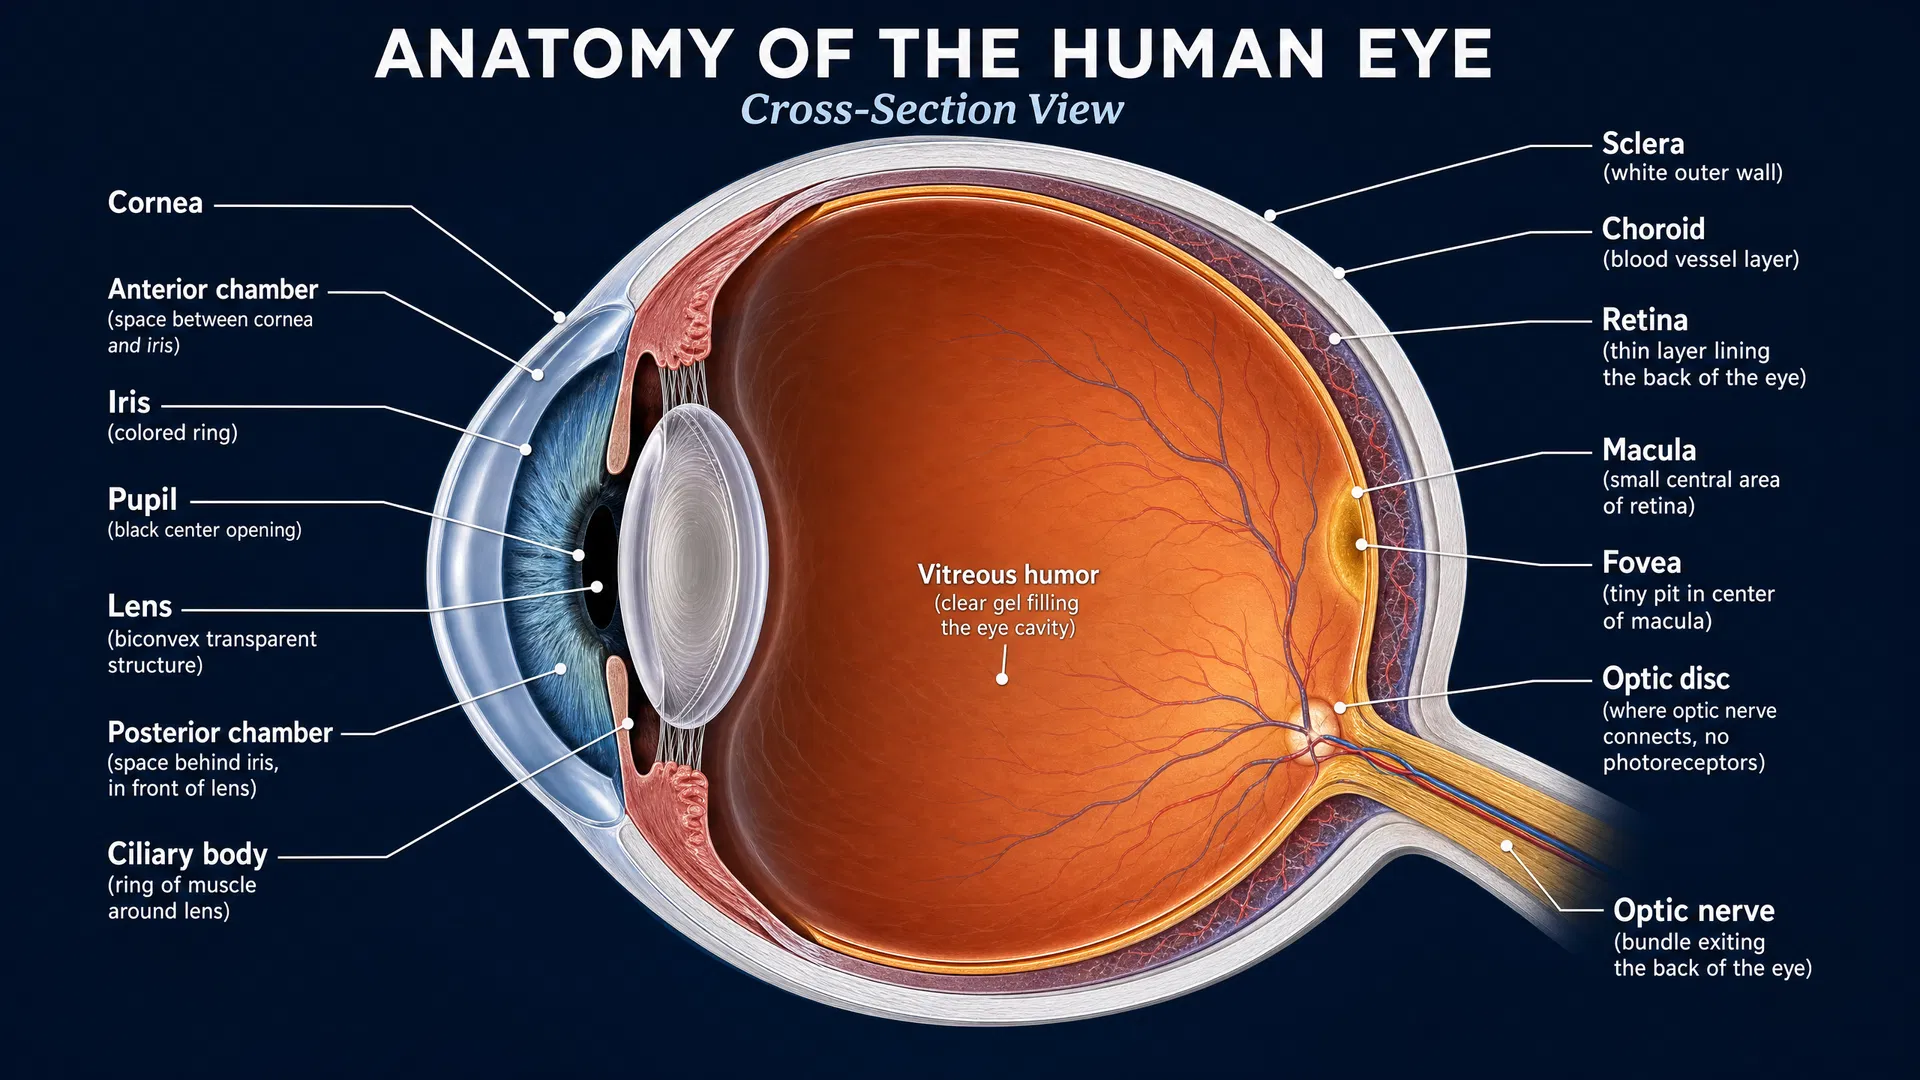
Stickler Syndrome, however, is primarily a structural disorder caused by mutations in collagen genes (such as COL2A1 and COL11A1). Many of these mutations exert a "dominant-negative" effect. This means the mutated gene produces an abnormal collagen protein that actively interferes with the healthy collagen produced by the normal gene copy, resulting in a structurally weak vitreous gel in the eye. Simply adding a healthy gene is not enough; the production of the toxic, abnormal protein must be stopped.
CRISPR and Allele-Specific Gene Editing
To overcome the dominant-negative hurdle, researchers are looking toward advanced gene-editing technologies like CRISPR-Cas9. Rather than just adding a gene, CRISPR acts as molecular scissors, capable of targeting and altering specific sequences of DNA.
In the context of Stickler Syndrome, a promising theoretical approach is "allele-specific knockout." The goal is to design a CRISPR system that specifically recognizes and disables the mutated copy of the collagen gene (the mutant allele) while leaving the healthy copy intact. If the mutant gene is silenced, the body would only produce healthy collagen from the remaining normal allele. While the total amount of collagen might be reduced, eliminating the interfering abnormal protein could allow for the formation of a much more stable vitreous structure, significantly reducing the risk of retinal detachment.
RNA-Based Therapies: Intercepting the Message
Another exciting avenue of research involves RNA-based therapies, specifically Antisense Oligonucleotides (ASOs). Before a gene is translated into a protein, its DNA sequence is transcribed into messenger RNA (mRNA). ASOs are small, synthetic pieces of genetic material designed to bind to specific mRNA molecules.
For Stickler Syndrome, ASOs could potentially be engineered to bind to the mRNA produced by the mutated collagen gene, either causing it to be degraded before the abnormal protein can be made or altering how the mRNA is spliced. This approach, known as "exon skipping," has shown promise in other genetic disorders like Duchenne muscular dystrophy and is being actively investigated for various inherited retinal diseases. Because ASOs do not permanently alter the patient's DNA, they offer a different safety profile compared to CRISPR, though they would likely require repeated administration.
The Road Ahead: From Lab to Clinic
It is important to emphasize that these advanced therapeutic approaches for Stickler Syndrome are currently in the preclinical phase. Researchers are actively developing cellular models and animal models that accurately mimic the human condition to test the safety and efficacy of these molecular tools.
The eye is an ideal target for these novel therapies because it is a small, enclosed compartment, allowing for localized delivery of treatments (via injection) with minimal systemic exposure. Furthermore, the eye is "immune-privileged," meaning it is less likely to mount a severe immune response to viral vectors or gene-editing tools.
Despite these advantages, significant challenges remain. Ensuring the precise delivery of gene-editing tools to the correct cells in the retina and vitreous, avoiding off-target genetic effects, and determining the optimal window for intervention (likely early in life before irreversible structural damage occurs) are all critical hurdles that must be cleared.
Conclusion
While a genetic cure for Stickler Syndrome is not yet available, the rapid pace of innovation in molecular biology provides substantial hope. As researchers continue to refine CRISPR technologies and RNA-based therapies, the prospect of treating the underlying cause of this complex connective tissue disorder is slowly moving from the realm of science fiction toward clinical reality.
Medical Disclaimer: This information is for educational purposes only and does not constitute medical advice. Genetic testing and clinical management should be performed by qualified healthcare professionals.