A Rare Dual Diagnosis in Two Brothers With X-Linked Retinoschisis
A 2026 case report in Frontiers in Genetics describes two brothers with both X-linked retinoschisis (XLRS), an inherited retinal condition, and X-linked ichthyosis (XLI), a skin disorder characterized by dry, scaly skin. According to the authors, this is the first reported family in which these two X-linked recessive conditions occurred together.
For families affected by XLRS, the report illustrates an important point: genetic conditions can coexist, and a careful assessment of both eye and non-eye symptoms can improve diagnosis. It also shows how genetic testing can clarify why family members may share some features while differing in others.
Two X-Linked Conditions, One Family
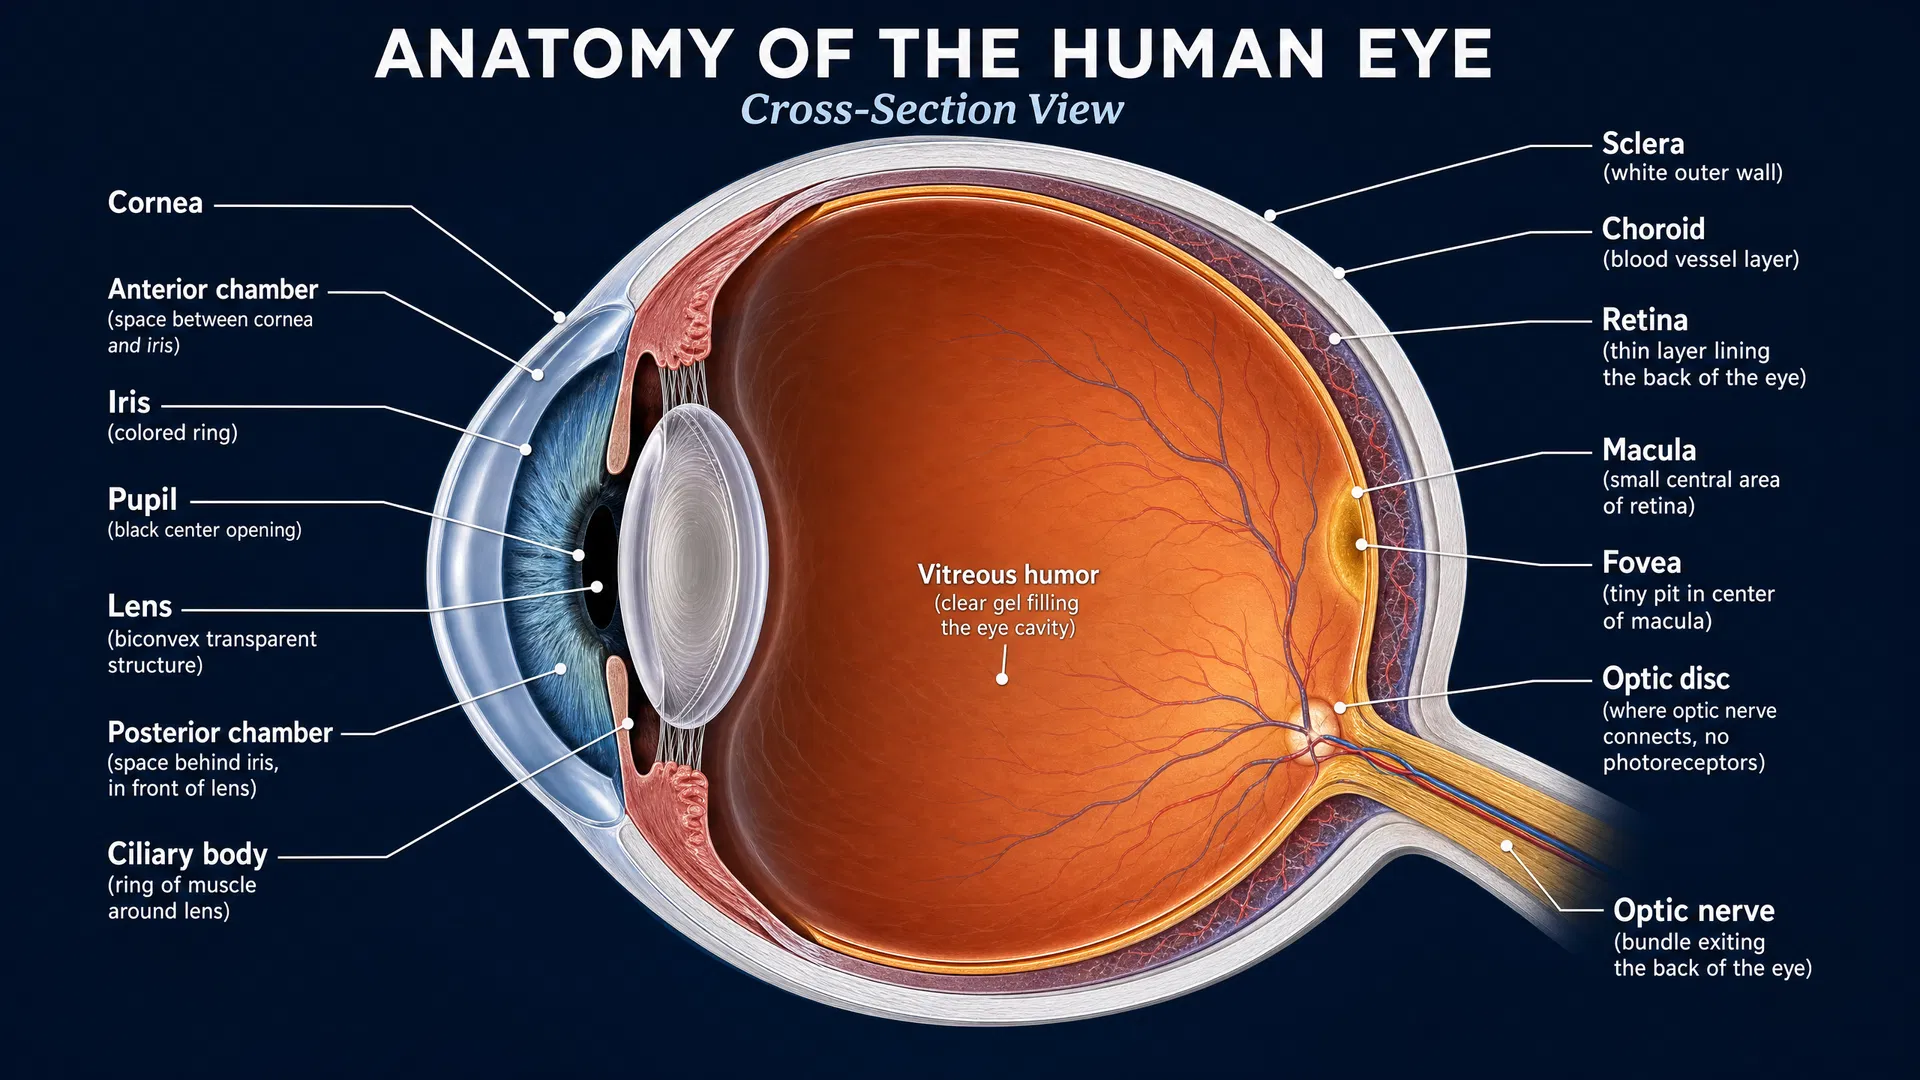
XLRS is caused by changes in the RS1 gene. In this family, both boys had reduced vision and characteristic retinal findings. Their eye examinations showed petaloid macular edema—a pattern of swelling in the central retina—and optical coherence tomography (OCT) revealed numerous cyst-like spaces in the macula of both eyes. These findings supported a diagnosis of bilateral XLRS.
The older brother was 11 years old and had decreased vision identified during a physical examination. His best-corrected visual acuity was 0.40 logMAR in each eye. His younger brother, age 6, had similar eye and systemic features. Neither boy had abnormalities in the front part of the eye, highlighting the value of retinal imaging when evaluating unexplained reductions in vision.
Both brothers also had dry, rough skin with diamond-shaped scales, a clinical feature consistent with XLI. Their skin symptoms had been noticed early: in the older brother, the parents recognized dry, rough skin one month after birth. The boys’ mother had previously been diagnosed with ichthyosis as well.
Genetic Testing Identified Changes in RS1 and STS
Genetic testing provided the explanation for the brothers’ combined presentation. Both carried the same RS1 variant: c.545G>T, resulting in p.Arg182Leu. The boys inherited this RS1 change from their mother. This finding linked their retinal findings to XLRS and adds to the reported mutation spectrum of XLRS in Chinese patients.
Testing also identified a deletion of approximately 478 kilobases on the X chromosome at Xp22.31. This deletion completely covered the STS gene region. Changes affecting STS are associated with XLI, explaining the brothers’ skin findings.
The report is especially notable because RS1 and STS are different genes with different roles and clinical consequences. RS1-related disease primarily affects the retina and vision, while STS-related disease is associated with the skin. In these brothers, the coexistence of both genetic findings matched the coexistence of both sets of symptoms.
Why Family Testing Matters
Because XLRS and XLI are inherited in an X-linked recessive pattern, family-based genetic evaluation was central to this case. The mother’s history of ichthyosis and the identification of the RS1 variant inherited from her helped establish the genetic context for her sons’ diagnoses.
The authors emphasize the phenotypic and genetic heterogeneity seen in inherited disease. Put simply, people in the same family may not present in exactly the same way, and a visible symptom in one body system may be relevant to an apparently separate concern in another. In this family, there was no reported family history of eye disease, yet both brothers had clear retinal manifestations on examination and OCT.
For families with XLRS, this underscores the value of sharing a complete medical and family history during an eye genetics evaluation. Skin findings, previous diagnoses, and results from relatives’ genetic tests may all help clinicians select and interpret testing.
Implications for Care and Future Treatment Research
This publication does not test a treatment, but it offers lessons that matter for treatment planning and research. First, precise molecular diagnosis can help ensure that retinal findings are attributed to the correct disease gene. In a family with more than one genetic condition, this distinction is essential for understanding which symptoms are related to XLRS and which are related to XLI.
Second, the case supports comprehensive clinical care. The boys required evaluation of both visual and skin manifestations rather than a single-condition approach. Retinal examination and OCT documented the eye effects of XLRS, while physical examination and testing clarified the skin diagnosis.
For future XLRS research, reports like this refine the catalog of RS1 variants seen in different populations. A more complete picture of genetic variation may improve diagnostic accuracy and support the design and interpretation of future gene-focused studies.
Looking Ahead
The first documented coexistence of XLRS and XLI in one family is a reminder that inherited retinal disease assessment is most powerful when clinical observations and genetic results are considered together. The brothers’ diagnoses were reached through the combination of vision testing, retinal imaging, skin examination, and family genetic analysis.
As inherited retinal disease research advances, detailed family reports will remain valuable. They help identify newly observed genetic combinations, expand knowledge of disease-associated variants, and encourage care teams to look beyond a single symptom or specialty. For people with XLRS and their families, this research reinforces that genetic answers can emerge from examining the whole person—and the whole family.